Parse counts, metadata, taxonomy, and phylogeny from a BIOM file.
Source:R/read_biom.r
read_biom.RdParse counts, metadata, taxonomy, and phylogeny from a BIOM file.
Arguments
- src
Input data as either a file path, URL, or JSON string. BIOM files can be formatted according to version 1.0 (JSON) or 2.1 (HDF5) specifications, or as classical tabular format. URLs must begin with
http://,https://,ftp://, orftps://. JSON strings are recognized by their first non-whitespace character being{. Compressed (gzip or bzip2) BIOM files are also supported. NOTE: to read HDF5 formatted BIOM files, theh5liteR package must be installed.- ...
Properties to set in the new rbiom object, for example,
metadata,id,comment, ortree. These override the corresponding content of the file.
Value
An rbiom object.
Details
The file's generated_by and date fields are kept in the returned object's
$generated_by and $date.
For classic tabular files, the first column must hold OTU IDs and a column
named taxonomy (or lineage) is parsed into taxonomic ranks. All other
columns are abundance counts: blank cells are read as 0 with a warning,
and columns containing non-numeric text are ignored with a warning.
Empty taxonomic ranks are NA.
Examples
    library(rbiom)

    infile <- system.file("extdata", "hmp50.bz2", package = "rbiom")
    biom <- read_biom(infile)

    print(biom)
#>
#> ══ Human Microbiome Project - 50 Sample Demo ═══════════════
#>
#> Oral, nasal, vaginal, and fecal samples from a diverse set
#> of healthy volunteers. Source: Human Microbiome Project
#> (<https://hmpdacc.org>).
#>
#>      50 Samples: HMP01, HMP02, HMP03, ..., and HMP50
#>     490 OTUs:    Unc01yki, Unc53100, LtbAci52, ...
#>       7 Ranks:   .otu, Kingdom, Phylum, ..., and Genus
#>       5 Fields:  .sample, Age, BMI, Body Site, and Sex
#>         Tree:    <present>
#>
#> ── 182 - 22k reads/sample ──────────────────── 2023-09-22 ──
#>

    # Taxa Abundances
    biom$counts[1:4,1:10] %>% as.matrix()
#>          HMP01 HMP02 HMP03 HMP04 HMP05 HMP06 HMP07 HMP08 HMP09 HMP10
#> Unc01yki     0     0     0     0     0     0     0     0     0     2
#> Unc53100  1083   543   301   223  2672   748  2436   321    57    28
#> LtbAci52     0     0     0     0     0     0     0     0     0     0
#> CnbTube3     0     0     0     0     0     0     0     0     0   659

    biom$taxonomy %>% head()
#> # A tibble: 6 × 7
#>   .otu     Kingdom  Phylum         Class          Order             Family Genus
#>   <chr>    <fct>    <fct>          <fct>          <fct>             <fct>  <fct>
#> 1 Unc01yki Bacteria Firmicutes     Bacilli        Lactobacillales   Lacto… Lact…
#> 2 Unc53100 Bacteria Firmicutes     Bacilli        Lactobacillales   Strep… Stre…
#> 3 LtbAci52 Bacteria Firmicutes     Bacilli        Lactobacillales   Lacto… Lact…
#> 4 CnbTube3 Bacteria Actinobacteria Actinobacteria Corynebacteriales Coryn… Cory…
#> 5 Unc02qsf Bacteria Firmicutes     Bacilli        Bacillales        Staph… Stap…
#> 6 PpbAcne6 Bacteria Actinobacteria Actinobacteria Propionibacteria… Propi… Prop…

    # Metadata
    biom$metadata %>% head()
#> # A tibble: 6 × 5
#>   .sample   Age   BMI `Body Site`   Sex
#>   <chr>   <dbl> <dbl> <fct>         <fct>
#> 1 HMP01      22    20 Buccal mucosa Female
#> 2 HMP02      24    23 Buccal mucosa Male
#> 3 HMP03      28    26 Saliva        Male
#> 4 HMP04      25    23 Saliva        Male
#> 5 HMP05      27    24 Buccal mucosa Female
#> 6 HMP06      32    25 Saliva        Male

    table(biom$metadata$Sex, biom$metadata$`Body Site`)
#>
#>          Anterior nares Buccal mucosa Mid vagina Saliva Stool
#>   Female              5             5         10      5     5
#>   Male                5             5          0      5     5

    sprintf("Mean age: %.1f", mean(biom$metadata$Age))
#> [1] "Mean age: 27.8"

    # Phylogenetic tree (plotting requires the 'ape' package)
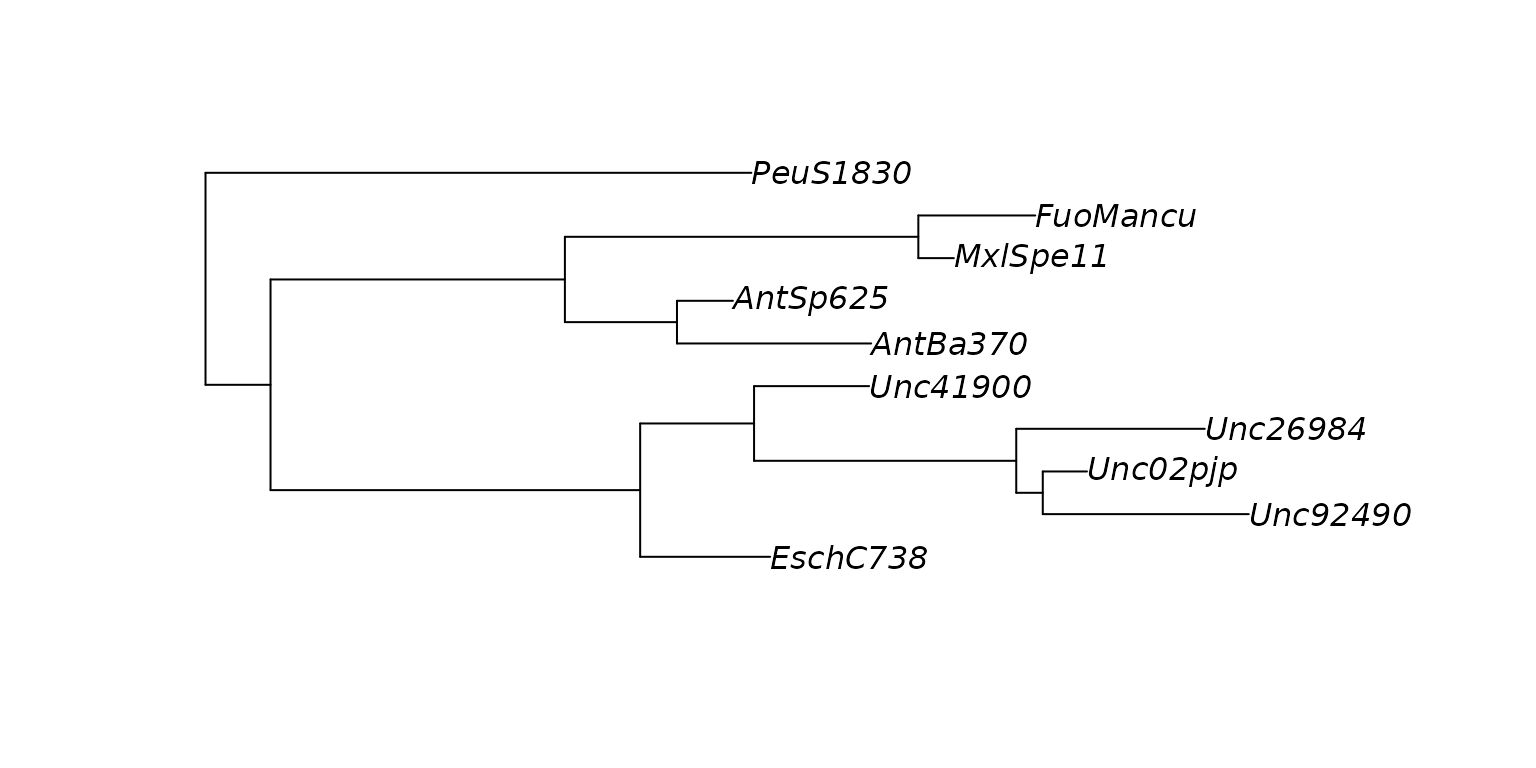
    tree <- tree_subset(biom$tree, 1:10)
    tree
#>
#> Phylogenetic tree with 10 tips and 9 internal nodes.
#>
#> Tip labels:
#>   EschC738, Unc92490, Unc02pjp, Unc26984, Unc41900, AntBa370, ...
#>
#> Rooted; includes branch length(s).
    if (requireNamespace("ape", quietly = TRUE))
      ape::plot.phylo(tree)